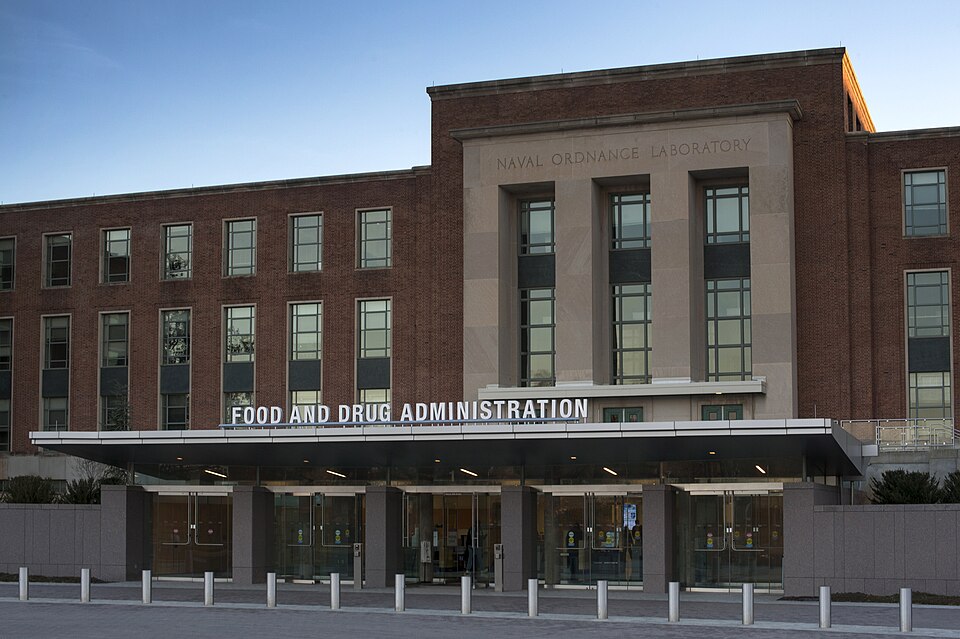
Photo: U.S. Food and Drug Administration (archive photo)
The Food and Drug Administration issued final guidance Monday calling on cell and gene therapy manufacturers to build potency assurance into every stage of development and production, replacing recommendations first published in 2011.
The FDA announcement addresses a basic question for some of medicine’s most complex products: does each batch retain the biological ability to do what the treatment is intended to do? The agency’s answer is that manufacturers should plan for that quality throughout the product’s life, rather than rely on a final test alone.
Cell and gene therapies are used or developed for serious conditions including cancers and rare diseases. Some are made for an individual patient, and some have short shelf lives. Those features make consistent production and timely quality checks especially important.
A broader strategy than the final laboratory test
The final potency-assurance guidance describes a science- and risk-based approach covering process design, manufacturing controls, raw materials, testing during production and assays used to release individual lots. An assay is a laboratory test used to measure a particular biological activity or characteristic.
Potency in this context is not simply the amount of material inside a vial. It concerns the product’s specific capacity to achieve its intended therapeutic effect. A batch can therefore require more than a basic count or concentration check to show that its active biological functions are being preserved.
FDA said its recommendations run from early investigational submissions through applications for a biologics license and continued verification after approval. The guidance also addresses viral-vector gene therapies, patient-specific treatments and products that cannot sit in storage while lengthy testing is completed.
Recommendations, not a new product approval
The final guidance sets out FDA’s recommendations for meeting existing quality expectations. It is not a new statute, does not itself approve any therapy and should not be read as a guarantee that a particular experimental treatment works. Guidance recommendations are distinct from binding legal requirements that apply to manufacturers.
For developers and production teams, the practical issue is how potency testing fits into a wider quality system. Controlling variability in raw materials and the manufacturing process can help reduce the risk of a weak or inconsistent batch before it reaches the point of final release. The agency describes its approach as flexible and based on a product’s risks, rather than a single test imposed on every therapy.
The document supersedes FDA’s January 2011 guidance, “Potency Tests for Cellular and Gene Therapy Products.” That change expands the discussion from testing toward a more complete assurance strategy across the product lifecycle.
FDA will hold a public webinar on December 10, from 1 p.m. to 2 p.m. Eastern. Registration is free but required and subject to capacity. The agency asks stakeholders to submit questions by November 18 and says questions will not be taken during the live event.
JBizNews Desk | Washington
© JBizNews.com. All rights reserved. Unauthorized reproduction or redistribution is strictly prohibited.